Privacy Disclosure Statement
Celexa
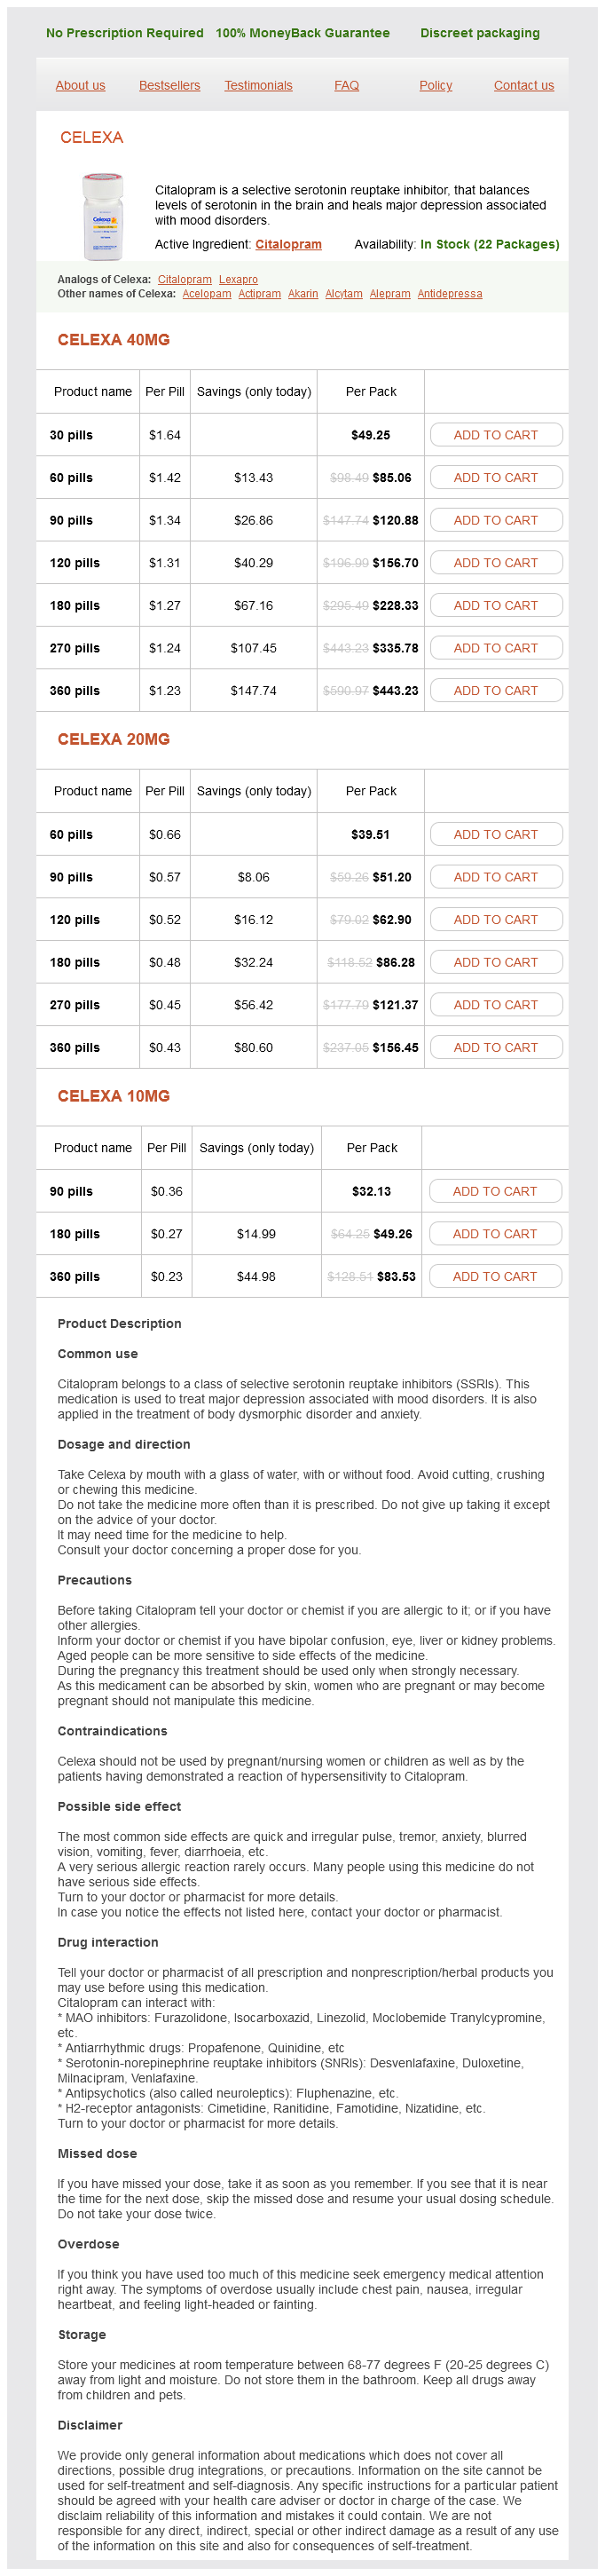
Celexa 40mg
- 30 pills - $49.25
- 60 pills - $85.06
- 90 pills - $120.88
- 120 pills - $156.70
- 180 pills - $228.33
- 270 pills - $335.78
- 360 pills - $443.23
Celexa 20mg
- 60 pills - $39.51
- 90 pills - $51.20
- 120 pills - $62.90
- 180 pills - $86.28
- 270 pills - $121.37
- 360 pills - $156.45
Celexa 10mg
- 90 pills - $32.13
- 180 pills - $49.26
- 360 pills - $83.53
Celexa dosages: 40 mg, 20 mg, 10 mg
Celexa packs: 30 pills, 60 pills, 90 pills, 120 pills, 180 pills, 270 pills, 360 pills
Only $0,25 per item
In stock: 807
Description
Pharmacogenetics-based versus conventional dosing of warfarin: a meta-analysis of randomized controlled trials treatment borderline personality disorder cheap 10 mg celexa mastercard. Aspirin plus clopidogrel as secondary prevention after stroke or transient ischemic assault: a systematic evaluate and metaanalysis symptoms 9 days post ovulation celexa 10 mg purchase line. Cardiovascular illness represents the number one explanation for death among adults in plenty of developed nations (Mozaffarian et al medicine vocabulary purchase celexa 20 mg line. The main lessons of lipoproteins and their properties are summarized in Table 33�2 treatment centers celexa 40 mg buy otc. Apoproteins have well-defined roles in plasma lipoprotein metabolism (Table 33�3). Mutations in lipoproteins or their receptors can lead to familial dyslipidemias and premature dying because of accelerated atherosclerosis. In all spherical lipoproteins, the most water-insoluble lipids (cholesteryl esters and triglycerides) are core components, and the extra polar, water-soluble parts (apoproteins, phospholipids, and unesterified cholesterol) are situated on the surface. Except for apo(a), the lipid-binding regions of all apoproteins comprise amphipathic helices that work together with the polar, hydrophilic lipids (such as floor phospholipids) and with the aqueous plasma environment by which the lipoproteins flow into. Differences in the non�lipid-binding areas determine the practical specificities of the apolipoproteins. Chylomicrons Chylomicrons are synthesized from the fatty acids of dietary triglycerides and cholesterol absorbed by epithelial cells in the small gut. In normolipidemic people, chylomicrons are current in plasma for 3�6 h after a fat-containing meal has been ingested. Apo B-48, synthesized only by intestinal epithelial cells, is unique to chylomicrons and capabilities primarily as a structural part of chylomicrons. Plasma Lipoprotein Metabolism Lipoproteins are macromolecular assemblies that comprise lipids and proteins. The lipid constituents embody free and esterified cholesterol, triglycerides, and phospholipids. The newly esterified and nonpolar ldl cholesterol moves into the core of the particle, which becomes progressively more spherical, larger, and fewer dense with continued ldl cholesterol acquisition and esterification. The transferred cholesterol subsequently is metabolized as a part of the lipoprotein into which it was transferred. Apo(a) of Lp(a) is structurally related to plasminogen and seems to be atherogenic. Atherosclerotic Cardiovascular Disease Risk Assessment Therapy for dyslipidemias is based on reducing the chance of deadly and nonfatal atherosclerotic cardiovascular events, including myocardial infarction and stroke. Secondary causes of dyslipidemias (Table 33�5), together with medicines that have an result on ldl cholesterol, should also be thought-about prior to initiating remedy. Cardiovascular danger assessment calculators have additionally been developed for lifetime risk of cardiovascular disease. Simvastatin and lovastatin are administered as inactive lactones that should be remodeled in the liver to their respective -hydroxy acids, simvastatin acid, and lovastatin acid. National lipid affiliation recommendations for patient-centered management of dyslipidemia: half 1-full report. Under steady-state circumstances, small quantities of the father or mother drug and its metabolites produced within the liver can be found in the systemic circulation. In the plasma, greater than 95% of statins and their metabolites are protein certain, aside from pravastatin and its metabolites, that are solely 50% bound. The t1/2 of the mother or father compounds are 1�4 h, besides in the case of atorvastatin and rosuvastatin, which have half-lives of about 20 h, and simvastatin with a t1/2 of about 12 h. The longer t1/2 of atorvastatin and rosuvastatin could contribute to their larger cholesterol-lowering efficacy. The liver biotransformsallstatins,andmorethan70%ofstatinmetabolitesare excreted by the liver, with subsequent elimination within the feces. Thus, statins with t1/2 of four h or less (all however atorvastatin and rosuvastatin) should be taken in the evening. Atorvastatin and rosuvastatin each have longer half-lives and could additionally be taken at different occasions of day to optimize adherence. Adverse Effects and Drug Interactions Hepatotoxicity Serious hepatotoxicity is rare and unpredictable, with a rate of about 1 case per million person-years of use. Myopathy refers to a broad spectrum of muscle complaints, starting from delicate muscle soreness or weak spot (myalgia) to life-threatening rhabdomyolysis. The danger of muscle opposed effects increases in proportion to statin dose and plasma concentrations. Consequently, elements inhibiting statin catabolism are associated with increased myopathy risk, including advanced age (especially > 80 years of age), hepatic or renal dysfunction, perioperative intervals, small body measurement, and untreated hypothyroidism. Concomitant use of drugs that diminish statin catabolism or interfere with hepatic uptake is related to increased danger of myopathy and rhabdomyolysis. The most typical statin interactions occur with fibrates, particularly gemfibrozil (38%), and with cyclosporine (4%), digoxin (5%), warfarin (4%), macrolide antibiotics (3%), and azole antifungals (1%). Coadministration of gemfibrozil nearly doubles the plasma focus of the statin hydroxy acids. When statins are administered with niacin, the myopathy probably is caused by an enhanced inhibition of skeletal muscle ldl cholesterol synthesis (a pharmacodynamic interaction). These pharmacokinetic interactions are related to elevated plasma concentrations of statins and their lively metabolites. However, the benefits of mixed remedy with any statin must be carefully weighed in opposition to the chance of myopathy. The choice of statins should be patient specific and based mostly on elements similar to cost, drug interactions, potential opposed effects, and desired depth. Rosuvastatin and pravastatin may be better tolerated than different statins and ought to be thought-about in patients with a history of myalgias with other statins. Lovastatin absorption is elevated when taken with food, and sufferers must be inspired to take with their evening meal. No more than 10 mg of simvastatin ought to be used in combination with diltiazem or verapamil. In distinction, different studies suggested statins may have a role within the prevention of dementias. Statins, especially at greater doses, doubtless confer a small elevated danger of growing diabetes. The bile acid sequestrants are highly positively Bold type signifies statins and doses used in randomized managed trials demonstrating a reduction in main cardiovascular occasions or dying. Because greater than 95% of bile acids are usually reabsorbed, interruption of this course of depletes the pool of bile acids, and hepatic bile acid synthesis will increase. Inhibition of reductase activity by a statin substantially will increase the effectiveness of the resins. The resin-induced improve in bile acid production is accompanied by an increase in hepatic triglyceride synthesis, which is of consequence in patients with important hypertriglyceridemia (baseline triglyceride degree > 250 mg/dL). Atorvastatin, lovastatin, and simvastatin are indicated for children eleven years and older. Statins are contraindicated during pregnancy and should be discontinued prior to conception if possible. Data concerning statin use while breastfeeding are limited, and use must be discouraged. In patients with normal triglyceride ranges, triglycerides could improve transiently after which return to baseline. When used with a statin, resins are often prescribed at submaximal doses due to poor tolerability. The powdered forms of cholestyramine (4 g/ dose) and colestipol (5 g/dose) are either blended with a fluid (water or juice) and drunk as a slurry or combined with crushed ice in a blender. Ideally, patients ought to take the resins earlier than breakfast and earlier than supper, starting with 1 scoop or packet twice day by day and growing the dosage after several weeks or longer as needed and as tolerated. Thestartingdoseis either 3 tablets taken twice daily with meals or all 6 tablets taken with a meal. Severe hypertriglyceridemia is a contraindication to the usage of cholestyramine and colestipol as a result of these resins improve triglyceride ranges. At current, there are insufficient information on the effect of colesevelam on triglyceride levels. These signs may be considerably decreased if the drug is totally suspended in liquid a number of hours earlier than ingestion. Constipation may occur however sometimes could be prevented by adequate every day water intake and psyllium. Colesevelam may be less doubtless than colestipol to trigger the dyspepsia, bloating, and constipation.
Zi hua di ding (Sweet Violet). Celexa.
- Are there safety concerns?
- What is Sweet Violet?
- Asthma, bronchitis, colds, congestion, cough, depression, flu symptoms, sleeplessness, lung problems, menopausal symptoms, nervousness, stomach problems, urinary problems, and other conditions.
- How does Sweet Violet work?
- Dosing considerations for Sweet Violet.
Source: http://www.rxlist.com/script/main/art.asp?articlekey=96245
In distinction treatment 4 autism celexa 20 mg generic overnight delivery, nonselective vasodilators such as adenosine or dipyridamole (which inhibits adenosine transmembrane transport and thereby increases extracellular concentrations) can worsen the perfusion of ischemic areas by dilating the comparatively constricted arterioles of the wholesome myocardium bad medicine 1 celexa 40 mg buy line, resulting in symptoms type 2 diabetes order celexa 20 mg with amex redistribution of blood move away from the ischemic myocardium ("steal phenomenon") symptoms electrolyte imbalance buy discount celexa 20 mg on line. In sufferers with angina owing to coronary artery spasm, the capacity of nitrovasodilators to dilate epicardial coronary arteries, significantly areas affected by spasm, is the primary mechanism of their beneficial effect. The impact could also be transient and incomplete in vivo, but abnormal "spasm" regularly is lowered. Indeed, many incidences of atypical chest ache and "angina" are due to biliary or esophageal spasm, and these also can be relieved by nitrates. Nitrates can even relax ureteral and uterine clean muscle, however these responses are of unsure medical significance. Multiple mechanisms have been proposed to account for nitrate tolerance, together with volume expansion, neurohumoral activation, mobile depletion of sulfhydryl teams, and the era of free radicals (Parker and Parker, 1998). Unfortunately, makes an attempt to prevent nitrate tolerance based mostly on these mechanisms. A clinically essential lesson of analysis on nitrate tolerance is that prolonged remedy with nitrates may not only induce a loss of response to nitrates, but in addition truly decrease angina threshold in the interval (Parker et al. If protection is inadequate, staff could experience severe headaches, dizziness, and postural weak spot during the first several days of employment ("Monday illness"). It appears prudent to not withdraw nitrates abruptly from a affected person who has received such remedy chronically. High doses must be prevented and therapy interrupted for 8�12 h daily, which allows the return of efficacy. Patients whose anginal sample suggests its precipitation by elevated left ventricular filling pressures. Toxicity and Untoward Responses Untoward responses to the therapeutic use of organic nitrates are virtually all secondary to actions on the cardiovascular system. Headache is frequent and may be severe, often reducing over a couple of days if therapy is sustained and often managed by decreasing the dose. Transient episodes of dizziness, weak point, and other manifestations associated with postural hypotension may develop, notably if the patient is standing motionless, and may progress occasionally to lack of consciousness, a response that seems to be accentuated by alcohol. It additionally could additionally be seen with very low doses of nitrates in patients with autonomic dysfunction. Even in extreme nitrate syncope, positioning and other measures that facilitate venous return are the only therapeutic measures required. In addition to visual disturbances, sudden one-sided listening to loss has also been reported. In the occasion that sufferers develop vital hypotension following mixed administration of sildenafil and a nitrate, fluids and adrenergic receptor agonists, if wanted, may be used for assist. Thus, many males wanting therapy for erectile dysfunction already may be receiving (or may require, particularly in the event that they improve physical activity) antianginal therapy. Headache, flushing, and rhinitis could also be observed, in addition to dyspepsia owing to rest of the decrease esophageal sphincter. Conditions corresponding to hypertension, anemia, thyrotoxicosis, obesity, heart failure, cardiac arrhythmias, and acute emotional stress can precipitate anginal signs in lots of sufferers. Table 27�1 lists the preparations and dosages of the nitrites and natural nitrates. The rapidity of onset, the length of motion, and the chance of growing tolerance are associated to the strategy of administration. The onset of motion is inside 1�2 min (fastest with the spray), and the results are undetectable by 1 h after administration. Anginal pain may be prevented when the drugs are used prophylactically instantly prior to exercise or stress. Godfraind and associates confirmed that the effect of the diphenylpiperazine analogues in stopping agonist-induced vascular clean muscle contraction might be overcome by elevating the focus of Ca2+ in the extracellular medium (Godfraind et al. In 1967, Fleckenstein advised that the unfavorable inotropic effect resulted from inhibition of excitation-contraction coupling and that the mechanism involved decreased motion of Ca2+ into cardiac myocytes. However, in variant angina, coronary constriction leads to decreased blood circulate and ischemic ache. Multiple mechanisms have been proposed to initiate vasospasm, together with endothelial cell injury. Whereas long-acting nitrates alone are often efficacious in abolishing episodes of variant angina, extra therapy with Ca2+ channel blockers normally is required. The utility of nitrovasodilators to relieve pulmonary congestion and to improve cardiac output in congestive heart failure is addressed in Chapter 28. In each smooth muscle and cardiac myocytes, Ca2+ is a set off for contraction, albeit by completely different mechanisms. Ca2+ channel antagonists, also called Ca2+ entry blockers or Ca2+ channel blockers, inhibit Ca2+ influx. In vascular smooth muscle, this leads to leisure, particularly in arterial beds, in cardiac myocytes to negative inotropic effects. All Ca2+ channel blockers exert these two principal actions, but the ratio differs based on the class as does the presence of chronotropic and dromotropic results. Clinically used Ca2+ channel blockers include the phenylalkylamine verapamil, the benzothiazepine diltiazem, and quite a few dihydropyridines, together with amlodipine, clevidipine, felodipine, isradipine, lercanidine, nicardipine, nifedipine, nimodipine, and nisoldipine. The constructions and relative specificities of representative medicine are proven in Table 27�2. Although these drugs are commonly grouped collectively as "calcium channel blockers," there are basic variations among verapamil, diltiazem, and the dihydropyridines with respect to pharmacodynamics, drug interactions, and toxicities. An increased focus of cytosolic Ca2+ causes increased contraction in each cardiac and vascular smooth muscle cells. In cardiac myocytes, the entry of extracellular Ca2+ causes a bigger Ca2+ launch from intracellular stores (Ca2+-induced Ca2+ release) and thereby initiates the contraction twitch. Cytosolic Ca2+ concentrations could be increased by numerous contractile stimuli in vascular easy muscle cells. Many hormones and autocoids increase Ca2+ inflow through so-called receptor-operated channels, whereas will increase in external concentrations of K+ and depolarizing electrical stimuli enhance Ca2+ influx through voltage-gated, or "potential operated," channels. The Ca2+ channel blockers produce their results by binding to the 1 subunit of the L-type voltage-gated Ca2+ channels and lowering Ca2+ flux through the channel. The vascular and cardiac effects of a variety of the Ca2+ channel blockers are summarized within the next part and in Table 27�2. Voltage-gated channels comprise domains of homologous sequence which are organized in tandem within a single massive subunit. In addition to the most important channel-forming subunit (termed 1), Ca2+ channels comprise several different associated subunits (termed 2, and) (Schwartz, 1992). Voltage-gated Ca2+ channels have been divided into no much less than three subtypes based mostly on their conductances and sensitivities to voltage (Schwartz, 1992; Tsien et al. Only the L-type channel is sensitive to the dihydropyridine Ca2+ channel blockers. All approved Ca2+ channel blockers bind to the 1 subunit of the L-type Ca2+ channel, which is the principle pore-forming unit of the channel. This roughly 250,000-Da subunit is related to a disulfide-linked 2 subunit of about 140,000 Da and a smaller intracellular subunit. The 1 subunits share a common topology of 4 homologous domains, every of which consists of six putative transmembrane segments (S1�S6). At least three distinct mechanisms may be responsible for contraction of vascular clean muscle cells. First, voltagegated Ca2+ channels open in response to depolarization of the membrane, and extracellular Ca2+ moves down its electrochemical gradient into the cell. After closure of Ca2+ channels, a finite period of time is required before the channels can open again in response to a stimulus. Emptying of intracellular Ca2+ shops might set off further influx of extracellular Ca2+ (store-operated Ca2+ entry), but its relevance in smooth muscle is unresolved. Third, receptor-operated Ca2+ channels enable the entry of extracellular Ca2+ in response to receptor occupancy. The Ca2+-calmodulin complicated in flip activates myosin light-chain kinase, with resulting phosphorylation of the myosin light chain. Such phosphorylation promotes interplay between actin and myosin and leads to sustained contraction of smooth muscle. Ca2+ channel blockers inhibit the voltage-dependent Ca2+ channels in vascular easy muscle and decrease Ca2+ entry. All Ca2+ channel antagonists chill out arterial smooth muscle and thereby lower arterial resistance, blood stress, and cardiac afterload. Although experimentally massive conductance veins of pig seem similarly or even more sensitive to Ca2+ channel blockers than arteries (Magnon et al.
Specifications/Details
Such studies decide how people will metabolize a selected drug and treatment 2nd degree burn buy celexa 40 mg online, to a restricted extent symptoms inner ear infection celexa 20 mg discount free shipping, predict the speed of metabolism symptoms of strep order 10 mg celexa. Interactions become a problem when a number of drugs are concurrently administered treatment ingrown hair buy 10 mg celexa fast delivery, for instance, in elderly patients, who on a daily basis might take prescribed anti-inflammatory medicine, cholesterol-lowering medication, blood stress medications, a gastric acid suppressant, an anticoagulant, and numerous over-the-counter medications. Similar research can be carried out with part 2 enzymes and drug transporters to predict the metabolic destiny of a drug. Computer-based computational (in silico) prediction of drug metabolism is a prospect for the near future. However, the buildings, decided by X-ray analysis of crystals of enzyme-substrate complexes, are static, whereas enzymes are versatile; this vital distinction may be limiting. Determining the potential for a drug candidate to produce acute toxicity in preclinical research is important and routine in drug development. This is typically done by administering the drug candidate to rodents at escalating doses, often above the anticipated human therapeutic dose. For drug candidates proposed for chronic use in humans, similar to for decreasing serum triglycerides and cholesterol or for treatment of kind 2 diabetes, long-term carcinogenicity research are carried out in rodent fashions. Signs of toxicity are monitored and organ harm assessed by postmortem pathologies. A new technology of high-throughput screening for biomarkers of toxicity is being adopted for drug improvement using metabolomics. Metabolomics is the systematic identification and quantification of all metabolites in a given organism or organic pattern. Using metabolomics, animals, either handled or not treated with a drug candidate, may be analyzed for the presence of a number of metabolites in urine that correlate with drug efficacy or toxicity. Metabolic fingerprints of particular compounds that are elevated in urine can be used to decide, in dose escalation research, whether or not a specific drug causes toxicity and can also be employed in early medical trials to monitor for potential toxicities. Metabolomics can be used to discover biomarkers for drug efficacy and toxicity that can be of worth in medical trials to establish responders and nonresponders. Drug metabolism could be studied in whole-animal mannequin systems and in people to decide the metabolites of a drug or point out the presence of a polymorphism in drug metabolism which may sign an antagonistic clinical consequence. Finally, biomarkers developed from experimental metabolomics could finally be developed for routine monitoring for signs of toxicity in sufferers receiving pharmacotherapy. Molecular clocks and the human condition: approaching their characterization in human physiology and illness. Inhibition of soluble epoxide hydrolase modulates inflammation and autophagy in obese adipose tissue and liver. Structural plasticity in the human cytosolic sulfotransferase dimer and its function in substrate selectivity and catalysis. Common sources of such variability embody noncompliance, medication errors, drug interactions (see Chapter four and Appendix I), and genetic factors. Pharmacogenomics, then again, research larger numbers of variants, in a person or throughout a inhabitants, to explain the genetic part of variable drug responses. Discovering which variants or combos of variants have useful penalties for drug effects, validating these discoveries, and finally applying them to patient care and to drug discovery are the duties of contemporary pharmacogenetics and pharmacogenomics. However, massive effect dimension single variants are the exception, and for many (most) drug responses, the genetic part of variable responses-although substantial-likely displays interacting influences of many genetic variants. A major problem to the sphere is to accrue giant numbers of topics with well-phenotyped drug responses to enable discovery, and subsequent replication and validation, of multigene results or of interactions of gene(s) with environmental factors. The nonfunctional alleles could be the same; the trait is then termed autosomal recessive, or different, during which case the subject is a compound heterozygote. In some cases, similar to clopidogrel, codeine, and irinotecan (described further on this chapter), variants in a single gene produce clearly outlined and clinically important differences in drug response. First, even within a single gene, a vast array of polymorphisms (promoter, coding, noncoding, completely inactivating, or modestly modifying) is possible. Each polymorphism could produce a special impact on gene perform and due to this fact differentially have an result on a measured trait. Second, even when the designations of recessive, codominant, and dominant are informative for a given gene, their utility in describing the genetic variability that underlies variability in drug response phenotype is diminished as a outcome of variability is commonly multigenic. Genetic variation may result in altered protein sequence and performance or in altered protein ranges by way of regulatory variation. Key genes involved in driving variable drug actions embrace these encoding drug-metabolizing enzymes, drug transport molecules, the molecular targets with which medicine interact, and a host of other genes that modulate the molecular context within which medicine act, notably genes dysregulated within the disease for which the drug is run. Drug metabolism is very heritable, as assessed utilizing drug exposures in monozygotic versus fraternal twins, drug exposures in cell strains from related subjects, or evaluation of very large information units utilizing technologies corresponding to genome-wide genotyping, mentioned additional in this chapter. Twin research advised that up to 75% of the variability in elimination half-lives for metabolized medicine may be heritable. Some drug metabolism traits behave in a standard "monogenic" style with three clearly definable (and separable) groups of drug response phenotypes: heterozygotes in addition to main and minor allele homozygotes. All of these mechanisms have been implicated in widespread germline pharmacogenetic polymorphisms. The term polymorphism was previously applied to variants occurring at a frequency higher than 1%. However, the appliance of genome sequencing to massive numbers of topics has made it clear that each individual has greater than 10 million sites across their genome at which they differ from some reference sequence. While a few of these are "widespread" (>1% frequency), the vast majority are a lot rarer. For uncommon variants clearly related to a genetic disease, the term mutation can also be used, but distinguishing between a really rare variant and a mutation could also be troublesome. There are 64 trinucleotide codons and solely 20 amino acids, so a number of codons encode the identical amino acid. The frequency of the *28 allele is up to 30%, with up to 10% of topics (depending on ancestry) being homozygous. For any gene, people could have two haplotypes, one maternal and one paternal in origin. A haplotype represents the constellation of variants that occur collectively for the gene on every chromosome. In some instances, this constellation of variants, quite than the individual variant or allele, may be functionally important. In others, however, a single variant could additionally be functionally essential regardless of other linked variants within the haplotype(s). With full linkage disequilibrium, genotype at one site is an ideal predictor of genotype on the linked web site. Patterns of linkage disequilibrium are inhabitants specific, and as recombination occurs, linkage disequilibrium between two alleles will decay and linkage equilibrium will end result. Ancestral Diversity Polymorphisms differ of their frequencies inside human populations and have been categorized as both cosmopolitan or inhabitants (or race and ethnic) particular. Cosmopolitan polymorphisms are these polymorphisms current in all ethnic groups and are more probably to be historic, having arisen before migrations of people from Africa, though present-day frequencies may differ amongst ancestral teams. The presence of ancestry-specific polymorphisms is according to geographical isolation of human populations. Individuals descended from a number of ancestries may show haplotype constructions and allele frequencies intermediate between their dad and mom. Pharmacogenetic Study Design Considerations There are many necessary concerns for the conduct of an experiment designed to establish sources of genetic variation contributing to variable drug responses. Pharmacogenetic Traits A pharmacogenetic trait is any measurable or discernible trait associated with a drug. Some traits reflect the useful or opposed effect of a drug in a affected person; decreasing of blood pressure or reduction in tumor size are examples. Other traits symbolize drug response "endophenotypes," measures which will more directly reflect the action of a drug in a biologic system and thus be extra amenable to genetic research but could additionally be faraway from the whole affected person or a whole population. Examples of the latter embody enzyme activity, drug or metabolite levels in plasma or urine, or drug-induced adjustments in gene expression patterns. A variant drug metabolizer phenotype could be inferred from genotype information or in some circumstances instantly measured by administering a "probe drug" (one thought to be metabolized by a single pathway) and measuring drug and metabolite concentrations. After assays are developed for a set of such variants, statistical strategies are used to relate genotype to phenotype. There are a number of databases that comprise info on polymorphisms in human genes (Table 7�1); these databases permit the investigator to search by gene for reported polymorphisms. Large-Scale "Agnostic" Approaches While the candidate gene approach has the intuitive attraction that recognized drug response pathways are studied, it has the disadvantage of trying only in regions of recognized biologic exercise.
Additional information:
Syndromes
- Blood in your urine, even once
- Antihistamines (if the condition is due to an allergy)
- Graves disease
- Parathyroid surgery, if the parathyroid levels become uncontrollably high
- Industrial accidents from falling asleep on the job
- Repair the aneurysm if the hemorrhage is due to an aneurysm rupture
- Overexposure to x-rays or other forms of radiation
- Cervical spondylosis with myelopathy (a problem with the vertebrae in the neck)
- Getting the area caught in a pant zipper
- Illness
Related Products
Usage: gtt.
Tags: discount celexa 10 mg visa, generic 10 mg celexa fast delivery, order 10 mg celexa visa, 20 mg celexa discount, celexa 40 mg generic line, cheap celexa 20 mg with visa, buy discount celexa 40 mg online, cheap celexa 10 mg overnight delivery, buy celexa 10 mg on-line, celexa 10 mg buy cheap, safe 40 mg celexa, celexa 10 mg order with amex, 10 mg celexa amex, generic 40 mg celexa with mastercard, order 10 mg celexa amex, cheap celexa 10 mg mastercard
10 of 10
Votes: 76 votes
Total customer reviews: 76
Customer Reviews
Pakwan, 61 years: Sevoflurane is a clear, colorless, unstable liquid at room temperature and should be saved in a sealed bottle.
Tjalf, 24 years: The causes include low ambient temperature, exposed physique cavities, chilly intravenous fluids, altered thermoregulatory control, and lowered metabolic rate.
Kapotth, 59 years: Ergot preparations should be administered as quickly as potential after the onset of a headache.
Raid, 28 years: Carbachol retains substantial nicotinic activity, significantly on autonomic ganglia.
Tyler, 22 years: Unlike schizophrenia, for which the biological understanding of monoamine neurotransmission has permitted synthesis of numerous effective compounds, no medication has yet been designed to treat the complete spectrum of bipolar dysfunction primarily based on biological hypotheses of the sickness.
Armon, 29 years: Acute administration of clonidine has been used within the differential diagnosis of patients with hypertension and suspected pheochromocytoma.
Inog, 53 years: The affinity of a competitive antagonist (Ki) for its receptor could be determined in radioligand binding assays or by measuring the functional response of a system to a drug in the presence of the antagonist (Cheng, 2004; Cheng and Prusoff, 1973; Limbird, 2005).
Altus, 49 years: A somewhat sordid historical past of fraud, battle of curiosity, and different irregularities has been revealed pertaining to the now-debunked affiliation research of thimerosal and autism; a long time of studies have been performed in safety reviews round this matter.